Is this for you?
Novel-device manufacturers where a suitable predicate may not exist and a risk-based De Novo classification could be relevant.
No matches.Browse services·Discuss your requirement
Medical Devices
Last reviewed: 2026-11-08
Considering an FDA De Novo route? Start with why a predicate may not fit, the device risk profile and available evidence. Tell us the current regulatory discussion so a specialist can separate De Novo from 510(k) or PMA planning.
Novel-device manufacturers where a suitable predicate may not exist and a risk-based De Novo classification could be relevant.
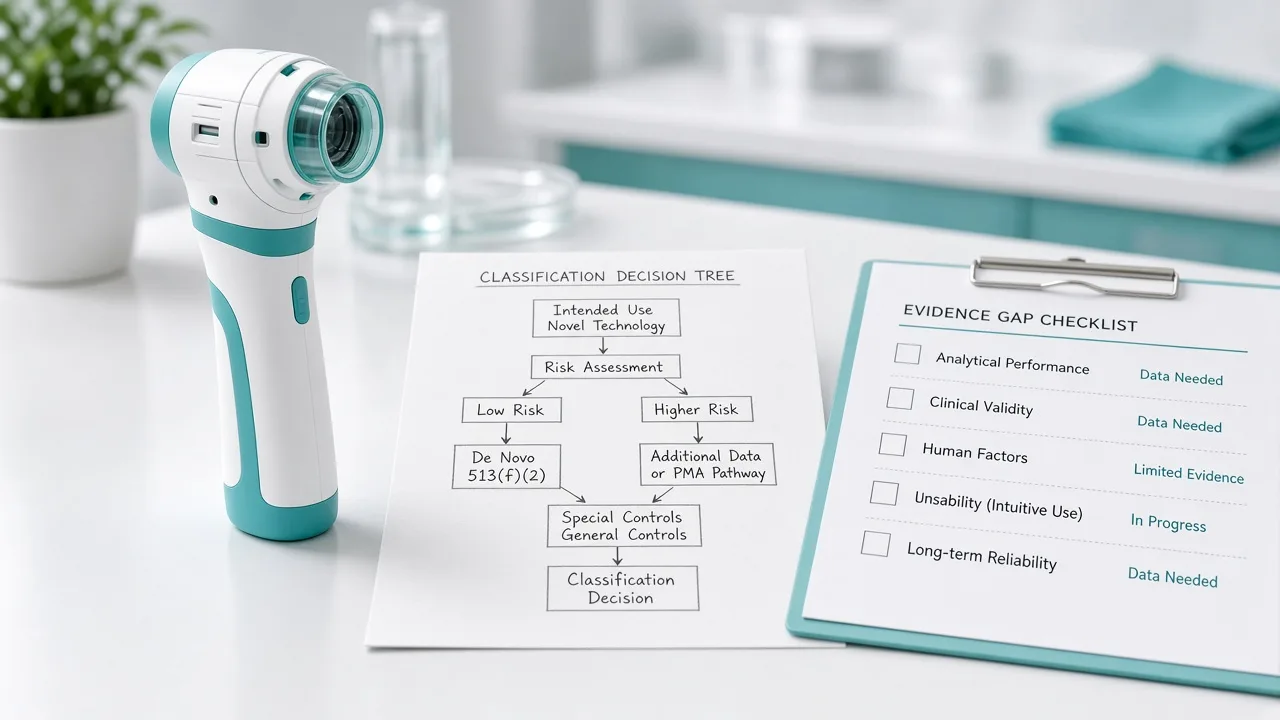
A novel low- to moderate-risk device without an appropriate predicate may warrant assessment of the FDA De Novo pathway. The route cannot be chosen from a marketing label alone. ConformArc can discuss an introduction to a provider that coordinates a specialist review of intended use, technology, classification history, risks, proposed controls and the evidence needed to support a request.
A useful early deliverable is a pathway decision memo that compares plausible alternatives, identifies missing performance and safety data, and sets out a submission work plan. Depending on the device, work may include bench, software, biological, electrical, usability or clinical evidence. Regulatory specialists should verify current FDA guidance and submission requirements. FDA decides whether to grant a De Novo request; ConformArc does not promise classification or market access.
No; eligibility and evidence need their own assessment.
Requirements can change. Confirm the current route and applicable edition for the specific product before starting the project.
Tell us the product or company requirement, target market, current stage and what your customer has asked for. We will review the initial details and reply about the next step. You can start without uploading confidential documents.
We use your details to respond to this enquiry. If a suitable external specialist could help, we will tell you who we propose to contact and ask before sharing your name, contact details or project information. The specialist may then contact you about this request. Read the Privacy Notice. See also how we choose providers.